
CAPA in Medical Devices: Fix the Problem, Not Just the Symptom.
What if the same quality problem keeps coming back even after you “fixed” it? That is usually a sign that something deeper has been missed. A problem gets identified, the immediate issue is corrected, everyone moves on. Then, a few weeks later, the same problem appeared again. This is exactly where CAPA became important.
CAPA is not a form to complete after something goes wrong. It's a way to really understand why a problem happened in the place. The focus is on finding the cause and making sure it doesn't happen again. In a medical device quality management system CAPA can begin for reasons. It might start with a customer complaint, an internal audit finding or a product that doesn't meet requirements. It could also come from a process failure, repeated problems, issues, with suppliers or trends seen in quality data. Every single sign tells us that something deeper must be examined. The goal is simple: learn from what went wrong and strengthen the system.
What is CAPA?
CAPA stands for Corrective Action and Preventive Action. At its core, CAPA asks two very practical questions:
- Why did this happen?
- What are we going to do so that it does not happen again?
A good CAPA process does not stop at fixing the immediate problem.
For example: Imagine a medical device manufacturer finds that a component has repeatedly failed during final inspection. The immediate reaction might be to replace the failed component. This solves the problem for that particular device. But it does not necessarily explain why the component keeps failing.
Questions arise:
- Was the supplier providing inconsistent material?
- Was the incoming inspection inadequate?
- Was the assembly process incorrect?
- Was the equipment improperly calibrated?
- Was the design itself contributing to the failure
CAPA pushes the organization to look beyond the obvious answer.
Correction vs Corrective Action
This distinction is one of the most important things to understand.
Correction: Correction deals with immediate problems.
Corrective Action: Corrective action deals with the cause of the problem.
Example A batch of devices fails a functional test.
Correction
The affected device is identified, repaired or rejected, and the immediate issue is contained.
Corrective Action
The organization investigates why the failure occurred, identifies the root cause, and changes the relevant process, procedure, training, supplier control, equipment, or other contributing factor.
The correction makes the problem go away today. While the Corrective Action is intended to stop it from becoming tomorrow's problem again.
Where do CAPAs come from?
CAPA does not always begin with a major failure. It can come from many sources, including:
- Customer complaints.
- Internal audits.
- External or Regulatory audits.
- Nonconforming products.
- Recurring manufacturing problems.
- Supplier issues.
- Process deviations.
- Risk management activities.
- Trend analysis.
- Post – market surveillance.
Sometimes one isolated issue may not require a CAPA. But when an organization starts seeing a pattern, that pattern deserves attention. This is why a good CAPA system looks at trends not just individual incidents.
What does a CAPA process look like?
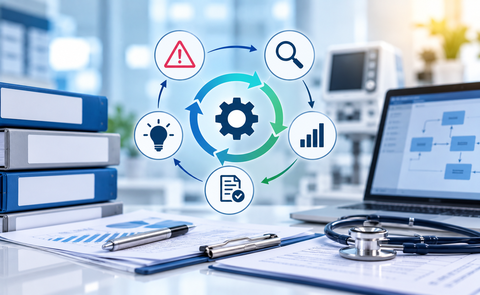
A practical CAPA process can be thought of a serious of questions;
What happened?
Clearly define the problem.
How serious is it?
Evaluate the impact and determine whether immediate containment is needed.
Why did it happen?
Investigate the root cause rather than stopping at the first obvious explanation.
What needs to change?
Define corrective and, where applicable, preventive action.
Did the action actually work?
Verify its effectiveness. Closing a CAPA does not simply mean completing an action item, the organization should have evidence that the action was effective.
A simple example
Let's say a manufacturer notices that several devices are failing a leakage test during final inspection. The team immediately retests the devices and repairs the ones that fail.
That's a correction.
But the quality team notices that similar failures have occurred several times over the last three months.
Now there is a trend.
The investigation discovers that the issue is related to an assembly step, and the existing work instruction does not clearly define the required assembly method.
The company then:
- Revise the work instruction.
- Trains the relevant operators.
- Reviews previously manufactured units where appropriate.
- Adds additional process controls.
- Monitors subsequent production batches.
After implementation, the company reviews the data to determine whether leakage failures have actually decreased.
That is the thinking behind an effective CAPA system.
Why is CAPA important?
A CAPA system gives an organization a way to learn from its problems.
Instead of treating every complaint, nonconformity, or audit finding as an isolated event, CAPA helps connect the dots.
A recurring problem can reveal a weak process.
A supplier issue can reveal a gap in supplier controls.
A complaint trend can reveal a product or design concern.
An audit finding can reveal that a procedure exists on paper but is not working effectively in practice.
In this way, CAPA can become a tool for continuous improvement, not just compliance. CAPA should not become a paperwork exercise.
One of the biggest mistakes is treating CAPA as:
Problem → Fill CAPA form → Add action → Close CAPA.
That may complete the paperwork, but it does not necessarily solve the problem.
A strong CAPA asks whether the organization has actually addressed the root cause and whether the solution is working.
The goal is not to have fewer open CAPAs. The goal is to have fewer recurring problems.
Final Takeaway
CAPA is ultimately about learning from what went wrong. A good CAPA does not ask only:
“How do we fix this?”
**It asks:
“Why did this happen, and what needs to change so that we don't keep fixing the same problem?”
Because in a medical device QMS, the best CAPA is not the one that closes the fastest.
**What about Preventive Action?
If corrective action asks:
“Why did this problem happen, and how do we stop it from happening again?”
Prevention action asks: “What could go wrong and what can we do before it becomes a problem?”
This is the key difference.
Corrective Action = Problem has happened.
For example, a manufacturer notices that a particular component is repeatedly failing during final inspection.
The organization investigates the root cause and changes the process to prevent the same problem from happening again.
That's corrective action.
Preventive Action = Problem hasn't happened yet.
Now imagine the quality team reviews supplier data and notices that a particular component's performance is gradually moving outside its normal range.
There has not been a product failure yet. But the trend suggests that a failure could happen in the future. The organization investigates the potential risk and takes action for example, tightening incoming inspection criteria, working with the supplier, or strengthening process controls. That's preventive action.
The simplest way to remember it:
Corrective Action:
Something went wrong → find the cause → prevent recurrence.
Preventive Action:
Something could go wrong → identify the potential cause → prevent occurrence.
And this is why a good QMS should not wait for a complaint, audit finding, or product before taking action. Sometimes the best quality problem is the one you prevent from happening in the first place.
One important point
Preventive action should not mean making changes simply because something might happen. There should be a reasonable basis for the action such as quality data, trends, audit results, risk information, supplier performance, complaints or other relevant information.
The goal is to use information proactively rather than waiting for a failure to provide the lesson. CAPA is therefore not just about fixing yesterday's problem. It is also about using what you know today to reduce the chance of tomorrow's problem.
V Reg Solutions helps medical device companies strengthen their QMS, implement effective CAPA processes, and maintain regulatory compliance.